How to Choose an OEM FIBC Pharmaceutical Supplier?

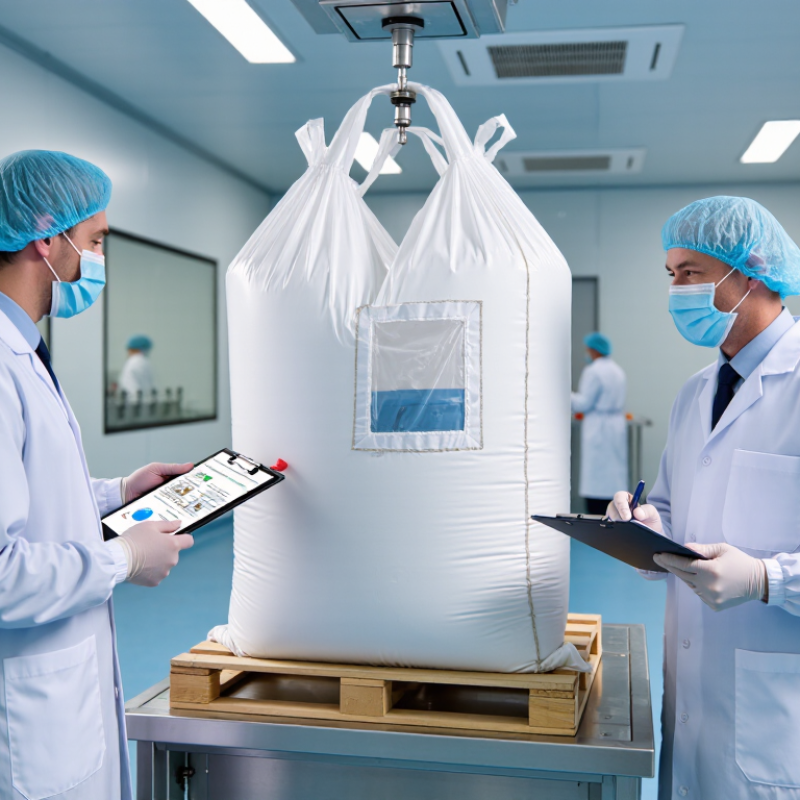
Choosing an Oem Fibc For Pharmaceutical Supplier requires more than comparing prices and printed specifications. Pharmaceutical bulk bags carry sensitive powders, so every material choice matters. The fabric, liner, seams, filling ports, and discharge system must support product protection and controlled handling.

In practice, a dependable supplier should explain its manufacturing process clearly. Ask about cleanroom controls, raw material traceability, electrostatic performance, and contamination prevention. Request batch records, certificates of analysis, and test results that match your intended application. A qualified supplier should also support sampling, technical drawings, and documented change control. Small details matter here. A loose seam can create dust. An unclear liner specification can affect product integrity.

Experience helps reveal weaknesses that brochures often hide. Visit the facility when possible, or request a remote quality audit with evidence. Review how the supplier manages complaints, deviations, training, and corrective actions. Relevant quality systems may include GMP-aligned practices and recognized ISO standards, depending on your market and product requirements. However, certification alone is not enough. It must reflect daily behavior.

The right Oem Fibc For Pharmaceutical Supplier should communicate openly and admit limitations. That point deserves attention. No supplier is perfect, and vague answers can become expensive problems later. Compare technical capability, documentation quality, delivery reliability, and after-sales support together. A careful evaluation protects the product, the operators, and your reputation. That is the real purpose of supplier selection.

How to Choose an OEM FIBC Pharmaceutical Supplier?

Define FIBC Needs: ISO 21898, GMP, and Pharmaceutical Contact Standards

When choosing an OEM FIBC pharmaceutical supplier, define the bag’s intended use before comparing prices. ISO 21898 provides a foundation for FIBC design, testing, handling, and safe filling. It does not prove pharmaceutical suitability by itself. Specify the working load, safety factor, filling method, discharge design, liner structure, and lifting configuration. A 1,000-kilogram bag may still fail if its outlet does not fit your equipment.

Product-contact materials need closer review. Request resin specifications, product-contact declarations, and evidence of controlled manufacturing. Depending on the medicine, assess extractables, leachables, particulate control, bioburden, and endotoxin risk. The supplier should explain whether the liner, coating, thread, ink, and sewing aids touch the product. Small details matter. A liner may look clean but release particles after vibration or heat exposure.

GMP principles should appear in daily operations, not only in a certificate folder. Check batch records, operator training, environmental controls, cleaning methods, and lot traceability. Ask how deviations, complaints, and material changes are managed. Require samples from a production-equivalent process, then test them under realistic filling and transport conditions. A common mistake is approving a sample made by hand. It may not represent routine output. Specifications can also become too broad, leaving quality teams unable to reject inconsistent bags. Review supplier audits, certificates of analysis, change notifications, and retention samples carefully. The documentation should be clear enough for a quality specialist to follow each bag back to its raw materials and production lot.

Audit Quality Systems Against 21 CFR Parts 210–211 and EU GMP

How to Choose an OEM FIBC Pharmaceutical Supplier?

An OEM FIBC supplier must withstand a quality audit, not just provide a clean sample. Check its written procedures against 21 CFR Parts 210–211. Part 211 contains 11 GMP subparts, covering personnel, facilities, production, laboratory controls, and records. Ask for controlled procedures, revision histories, training records, and deviation investigations. Small gaps matter.

The FDA’s 2023 Quality Metrics Annual Report highlights three practical indicators: lot acceptance rate, product quality complaint rate, and invalidated out-of-specification results. Request these metrics for the FIBC production line, where available. Compare complaint trends over three years. A polished audit room proves little. Review batch records, line-clearance checks, cleaning validation, and change controls. Confirm that liner materials, stitching threads, and contact surfaces have defined specifications. Sampling plans should be risk-based and traceable to each lot.

EU GMP Volume 4 uses nine core chapters, including documentation, production, quality control, and outsourced activities. The supplier should map these expectations to its own system. Verify supplier qualification, subcontractor oversight, and transport controls. Ask how it manages fiber shedding, electrostatic risks, moisture exposure, and seal integrity. Test evidence should include methods, acceptance limits, and laboratory approval status. One uncomfortable point remains: many audits overvalue certificates and undervalue records. I would not accept that shortcut. A supplier may pass a visit while weak change control remains hidden. Insist on recent examples.

Verify Materials with USP <661.1>, USP <665>, and BPOG Testing

Choosing an OEM FIBC pharmaceutical supplier requires more than checking a cleanroom certificate. Ask how the supplier qualifies polymers, liners, coatings, threads, and processing aids. USP <661.1> helps assess plastic materials of construction, including chemical composition and biological safety. Request the exact resin grade, additives, test conditions, and current reports. Generic declarations are weak evidence.

USP <665> focuses on plastic components and systems used during pharmaceutical manufacturing. It can help evaluate extractables from an FIBC liner, discharge port, or inner film. Confirm whether the test article matches your proposed bag design. A different film thickness can change results. It matters. BPOG testing adds practical extractables data under process-relevant conditions. Review solvents, temperatures, contact times, and analytical limits. Do not accept a report without its protocol.

A reliable OEM should connect every test result to a material lot and production record. Check change-control procedures, supplier qualification, cleaning validation, and packaging integrity tests. Ask for samples from routine production, not only development batches. One audit mistake is treating USP compliance as automatic product approval. It is not. Suitability depends on drug contact, process conditions, exposure time, and risk assessment. Some documentation may appear complete but still lack raw data. That gap deserves attention. The supplier should explain deviations clearly, preserve traceability, and repeat testing after meaningful material or design changes. A technically strong partner also admits uncertainty. That is often more trustworthy than perfect-looking paperwork.

Set Cleanliness Targets Using ISO 14644-1 and USP <788> Limits

How to Choose an OEM FIBC Pharmaceutical Supplier?

Set cleanliness targets before comparing OEM FIBC suppliers. ISO 14644-1 classifies airborne cleanliness by particle concentration in controlled rooms. It does not prove that an FIBC surface is sterile or particle-free. Ask for the filling-room classification, particle-size data, monitoring frequency, and recent deviation records. A reliable supplier should connect these records to the actual manufacturing step, not only display a certificate.

USP <788> addresses particulate matter in injections. Its limits commonly include particles equal to or larger than 10 µm and 25 µm, but requirements vary by product type and container volume. For example, small-volume injections have commonly used limits of 6,000 particles at ≥10 µm and 600 at ≥25 µm per container. Confirm the current compendial requirement for your product. Then define FIBC controls that support it, such as double-bagging, controlled unpacking, visual inspection, and validated particle testing. The FIBC is only one part of the contamination-control strategy.

Tips: Request raw particle-count reports, not only pass/fail statements. Check sampling locations and instrument calibration. Ask how operators handle liners, ports, and closures. In practice, supplier documents can look complete while missing a clear link to your process risk. That gap deserves attention. A practical audit should include a room walkthrough, material-flow review, and traceability check from resin or fabric to released FIBC. Avoid targets chosen only because they sound clean. They must be measurable, justified, and reviewed when your process changes.

How to Choose an OEM FIBC Pharmaceutical Supplier? - Set Cleanliness Targets Using ISO 14644-1 and USP <788> Limits
Qualification Dimension Reference Standard Recommended Requirement Acceptance Target or Limit Supplier Evidence to Request Why It Matters for Pharmaceutical FIBCs
Cleanroom Classification and Airborne Particle Control
Final FIBC assembly environment ISO 14644-1:2015 Document the ISO classification of the room where the cleaned liner, body, discharge components, and other product-contact parts are assembled. Define the required class by risk assessment; ISO Class 7 or ISO Class 8 may be used only when justified by the process and product-contact design. Current cleanroom classification report, room layout, sampling plan, and classification date. ISO 14644-1 classifies airborne particle concentration by particle size. It does not by itself prove sterility or microbial control.
ISO Class 5 airborne particles ISO 14644-1:2015 Use as a benchmark for critical open-product or exposed product-contact operations when the risk assessment requires a highly controlled zone. Maximum particles per cubic metre: 3,520 at ≥0.5 µm and 29 at ≥5.0 µm. Classification data at ≥0.5 µm and ≥5.0 µm, including occupancy state and sampling locations. Provides a stringent airborne cleanliness reference for high-risk exposure points, but it is not a direct FIBC product-release limit.
ISO Class 6 airborne particles ISO 14644-1:2015 Consider for controlled component preparation or assembly areas where a tighter environment is required than a conventional controlled room. Maximum particles per cubic metre: 35,200 at ≥0.5 µm and 293 at ≥5.0 µm. Routine monitoring records, alert/action levels, HEPA filter maintenance, and airflow verification. Helps reduce airborne particulate deposition during the preparation of liners and product-contact components.
ISO Class 7 airborne particles ISO 14644-1:2015 Use as a possible target for controlled FIBC component assembly when supported by a documented contamination-control strategy. Maximum particles per cubic metre: 352,000 at ≥0.5 µm and 2,930 at ≥5.0 µm. Room classification certificate, differential-pressure records, temperature and humidity records, and environmental-monitoring trend reports. Offers a practical controlled-environment benchmark for many pharmaceutical packaging operations.
ISO Class 8 airborne particles ISO 14644-1:2015 Use only where the product-contact risk assessment permits a less stringent controlled environment and the process has effective cleaning and inspection controls. Maximum particles per cubic metre: 3,520,000 at ≥0.5 µm and 29,300 at ≥5.0 µm. Classification report, cleaning validation or verification records, personnel-gowning procedure, and deviation history. May be suitable for lower-risk preparation steps, but it should not automatically be treated as adequate for every pharmaceutical application.
Particle-size reporting ISO 14644-1:2015 Require particle data at the specified ISO class sizes and ensure the sampling plan reflects the room area and operational state. At minimum, report ≥0.5 µm and ≥5.0 µm concentrations for the ISO class comparison. Calibrated particle-counter certificates, raw data, sample locations, room state, and calculation method. Incomplete particle-size reporting can make an ISO classification claim difficult to verify or compare.
USP <788> Particulate-Matter Considerations
Application of USP <788> USP <788> Particulate Matter in Injections Use USP <788> as a reference for the pharmaceutical product or solution that will be filled, not as a standalone cleanliness specification for an empty FIBC. Do not describe an FIBC as USP <788>-compliant unless the specific test sample, extraction method, and intended use have been defined and validated. Written applicability statement, product-contact risk assessment, extractables or rinse-test protocol, and laboratory test records where applicable. USP <788> establishes limits for particulate matter in injections. It does not classify manufacturing rooms or directly set limits for packaging materials.
Small-volume injection reference limit USP <788>, light-obscuration particle count method Use only when the filled pharmaceutical product is a small-volume injection and the monograph or applicable regulation requires this criterion. Not more than 6,000 particles per container at ≥10 µm and not more than 600 particles per container at ≥25 µm. Validated test method, sample preparation records, instrument suitability, and batch-level particulate results. Useful for defining the particulate-control outcome expected of the complete filling system, including packaging-related contamination risks.
Large-volume injection reference limit USP <788>, light-obscuration particle count method Use only when the filled pharmaceutical product is a large-volume injection and the applicable specification requires this criterion. Not more than 25 particles per mL at ≥10 µm and not more than 3 particles per mL at ≥25 µm. Validated sample volume, dilution or preparation instructions, instrument qualification, and investigation procedure for excursions. Converts the particulate expectation to a concentration-based limit appropriate for large-volume injectable products.
Visual inspection relationship USP <788> and applicable visual-inspection requirements Require documented visual inspection of FIBCs and product-contact components in addition to particle-count testing where appropriate. No visible foreign matter, fibers, loose threads, damaged seams, powder residue, or packaging debris according to the approved inspection specification. Inspection procedure, lighting requirements, inspector qualification, defect library, and inspection records. Particle-count testing may not detect every visible defect or fiber that could affect pharmaceutical processing.
OEM Supplier Qualification and Release Controls
Cleaning and component preparation Supplier quality system and contamination-control plan Require a documented, repeatable cleaning process for fabric, liners, closures, and other product-contact components. Approved cleaning method with defined parameters, equipment status, personnel training, and batch traceability. Cleaning SOP, validation or verification summary, equipment-maintenance records, and cleaning-batch records. Cleaning controls should prevent residues, loose fibers, dust, and cross-contamination before final assembly.
Packaging after cleaning Contamination-control plan Protect cleaned FIBCs immediately after inspection and seal them in a controlled manner suitable for transport and storage. Double-bagging or an equivalent validated protective method when required by the risk assessment; tamper-evident identification recommended. Packaging specification, seal-integrity checks, labeling procedure, storage conditions, and transport qualification where applicable. Even a clean FIBC can be recontaminated by handling, warehouse dust, moisture, or damaged outer packaging.
Microbiological control Risk-based pharmaceutical quality system Define bioburden or microbiological monitoring requirements separately from ISO 14644-1 particle classification. Set numerical limits through a documented risk assessment, product requirement, and validated test method; ISO 14644-1 does not provide microbial limits. Environmental microbiology program, bioburden method, alert/action limits, trend reports, and excursion investigations. Particle cleanliness and microbial cleanliness are different attributes and require separate controls and evidence.
Lot traceability Good manufacturing and supplier-quality practice Require traceability from raw material and cleaning batch through assembly, inspection, packaging, and shipment. Unique lot number, production date, material identification, operator or line record, and retained quality documentation. Certificate of conformance, batch record, raw-material certificates, inspection report, and change-control history. Traceability supports effective investigations, recalls, change assessment, and pharmaceutical customer audits.
Change control Supplier quality agreement Require prior notification and documented assessment for changes to resin, fabric, liner, cleaning agent, equipment, site, process, or packaging. No critical change implemented without customer review or approval where specified in the quality agreement. Change-control SOP, notification procedure, historical change log, and requalification plan. Uncontrolled changes can alter particulate shedding, extractables, cleanliness, mechanical performance, or compatibility.
Final release decision Approved specification and quality agreement Release each lot against agreed cleanliness, visual, dimensional, mechanical, and documentation requirements. All critical acceptance criteria met; deviations investigated and formally dispositioned before shipment. Certificate of analysis or conformance, inspection results, deviation report, and shipment-release authorization. A structured release decision converts cleanliness targets into consistent, auditable supplier performance.
Interpretation note: ISO 14644-1 limits above are airborne particle concentrations expressed as maximum particles per cubic metre for the stated particle sizes. USP <788> limits apply to particulate matter in injections and should not be treated as direct release limits for empty FIBCs without a justified, validated test strategy. Final requirements should be established through a documented pharmaceutical risk assessment and supplier quality agreement.

Compare Safety, Traceability, and 5:1 FIBC Load-Test Performance

Choosing an OEM FIBC pharmaceutical supplier requires more than a clean facility and attractive test certificates. Ask how the supplier controls resin, fabric, thread, liners, and stitching during every production batch. Safety begins with material compatibility, static-control planning, hygienic handling, and clearly defined filling and discharge procedures. Request documented risk assessments, inspection records, and corrective-action history. Small gaps matter. Traceability should connect each bag to raw-material lots, operators, equipment, inspection results, and release approval. A scannable batch code helps, but only when records remain complete and retrievable.

For a 5:1 FIBC, the rated safe working load represents one-fifth of the tested failure load under specified conditions. Do not accept “5:1” as a standalone promise. Review the test method, sample quantity, conditioning, load duration, deformation limits, and laboratory competence. Ask whether the tested design matches the final bag, including seams, lifting loops, liner, and closure. A strong report shows failures, not only successes. That detail builds trust. Test results should support your process validation, not replace it.

During supplier audits, observe actual line practices rather than relying on polished presentations. Check how workers prevent fiber contamination, segregate materials, and quarantine nonconforming bags.

I have seen traceability systems look impressive until a missing liner lot delayed an investigation. That weakness deserves honest discussion. A reliable OEM partner explains limitations, repeats critical tests, and updates documents after design changes. Still, no supplier can remove every operational risk. Your team must confirm compatibility, handling, storage, and release criteria before routine use.